Medical Device Recall Logistics: FDA Guide
Key takeaways
A medical device recall is a correction or removal of a device that violates FDA law, and FDA classifies it as Class I, II or III by relative health risk.
Under 21 CFR Part 806, manufacturers and importers must report a qualifying correction or removal to FDA within 10 working days of starting it.
Medical device recall logistics covers notification, retrieval, quarantine, disposition and replacement, all traceable to lot, serial number or UDI.
Cross-border recalls need a plan for each country's authority, export documents for returned units and local destruction where returning is not practical.
FDA expects records that show who received the device, who was notified, what came back and what happened to it, kept in a form that supports status reports and termination.
A medical device recall is as much a logistics operation as a regulatory one. Once a company decides to correct or remove a device, it has to find every affected unit, tell every consignee, bring product back or fix it in place, keep it segregated, decide what happens to it and, often, ship replacements at the same time. Medical device recall logistics is the work of doing all of that quickly and with a record of every step that FDA can review.
This guide explains how FDA medical device recalls work, the logistics tasks in each phase, how reverse logistics works for recalled devices, what changes when a recall crosses borders and which records FDA expects. It reflects the regulations as published in the eCFR in October 2026. Your regulatory team decides on classification, reporting and strategy. The logistics team makes that strategy happen.
What happens in a medical device recall?
In a medical device recall, the company identifies a problem, decides on a correction or removal, notifies FDA and its customers, carries out the action, checks that it reached everyone and asks FDA to terminate the recall once the product is accounted for. The FDA explains that a correction addresses a problem where the device is used or sold, while a removal takes the device away from that location. FDA uses the term recall when a correction or removal addresses a device that violates FDA law.
A recall does not always mean returning product. FDA notes that a recall sometimes means a device needs to be checked, adjusted or fixed, and that a recalled implant does not always have to be removed from patients. The logistics plan therefore depends on the action chosen. In broad terms, a recall moves through these phases:
Assessment. The company evaluates the problem, the health risk and which lots, models or serial numbers are affected.
Strategy. It sets the depth of the recall (wholesale, retail or user level), whether a public warning is needed and the level of effectiveness checks.
Notification. It notifies FDA and each direct account, with instructions on what to do with the product.
Execution. Product is held, corrected on site or retrieved, and replacements are shipped where needed.
Effectiveness checks. The company verifies that consignees received the notice and acted on it.
Disposition and termination. Returned product is reworked, relabeled or destroyed, and the company asks FDA to terminate the recall.
Under 21 CFR Part 7, Subpart C, a firm that recalls voluntarily need not wait for FDA to review its strategy before starting the removal or correction. That is why logistics should be ready on the day the decision is made.
What are the FDA medical device recall classes?
The FDA medical device recall classes are Class I, Class II and Class III, assigned by FDA after a health hazard evaluation to show the relative degree of risk. Class I is the most serious and Class III the least. The definitions below follow FDA's own wording.
Class | FDA definition | What it usually means for logistics planning |
|---|---|---|
Class I | A reasonable chance that a product will cause serious health problems or death | Highest priority, often user-level depth, fastest notification and retrieval, likely replacement shipments |
Class II | A product may cause a temporary or reversible health problem, or there is a slight chance of serious health problems or death | Planned retrieval or field correction, coordinated with sites and distributors |
Class III | A product is not likely to cause any health problem or injury | Structured removal or correction, often at wholesale or retail depth |
FDA does not set fixed logistics deadlines by class. Timelines come from the recall strategy, the health risk and FDA's instructions in each case. Whatever the class, 21 CFR 7.49 says recall letters and envelopes should be marked urgent for Class I and Class II recalls, and every class requires the same discipline in records. FDA publishes classified recalls in its weekly Enforcement Report.
The reporting clock under Part 806
21 CFR Part 806 requires device manufacturers and importers to report to FDA any correction or removal started to reduce a risk to health, or to remedy a violation that may present a risk to health, within 10 working days of starting it. If the same action is later extended to more lots, the report must be amended within 10 working days of the extension. Some actions are exempt, such as market withdrawals, routine servicing and stock recovery, but those still need internal records. The logistics team should know the start date, because it starts the reporting clock.
How is reverse logistics handled in a medical device recall?
Reverse logistics in a medical device recall is handled by building a retrieval list from distribution records, sending each site return instructions and packaging, collecting units with a documented chain of custody, receiving them into a segregated quarantine area, reconciling what came back against what was shipped, and recording the final disposition of every unit. Medical device reverse logistics differs from routine returns because every unit must be accounted for and every step must be provable.
Build the retrieval map
Start from shipment and sales records, down to the lot or serial number of each consignee. Hospital systems often move devices between their own sites, and distributors resell to secondary accounts, so the first list is rarely complete. Ask each direct account to confirm quantities on hand and to identify any onward customers, as the recall communication should request where appropriate.
Serialized tracking for recalls
Serialized tracking for recalls is what makes reconciliation possible. Part 806 defines the unique device identifier (UDI) as a device identifier plus a production identifier that can include the lot, the serial number, the expiration date and the manufacturing date. When labels carry a UDI, scan it at pickup, at receipt and at disposition. A single scan record per unit, linked to the consignee, is the simplest way to answer how much product is still in the field.
Pick up, quarantine and disposition
The table shows the reverse logistics steps and the record each one should produce.
Step | Logistics action | Record to keep |
|---|---|---|
Return kit | Send packaging, labels and instructions to each site | Kit shipment tracking and site contact |
Collection | Carrier, courier or field team picks up units | Pickup confirmation with quantities and identifiers |
Transit | Tracked transport, temperature control where the device requires it | Tracking events, temperature data, chain of custody |
Receipt | Scan into a segregated quarantine location | Receiving log reconciled to the consignee list |
Disposition | Rework, relabel, return to stock after correction, or destroy | Disposition decision, approval and destruction certificate |
Some device categories need temperature control on the way back, for example diagnostic products with reagent components, so that units arrive in a condition that supports investigation. Keep returned units physically separated from saleable stock and clearly labeled. Our articles on reverse logistics in life sciences and chain of custody in life science shipping cover the principles in more depth.
Shipping replacements
A recall often creates an outbound problem at the same time. A hospital that returns a critical device still needs one. For local sites, a same-day courier can collect the recalled unit and deliver the replacement in one visit. For distant sites, next flight out places the replacement on the earliest suitable commercial flight, and an onboard courier keeps a device in personal custody for the whole journey. For higher volumes, air freight to regional points followed by local delivery is usually more efficient. Each replacement shipment should reference the recall and the units it replaces, so the reconciliation stays clean.
How do you manage a recall that crosses international borders?
You manage a recall that crosses international borders by running a separate notification and logistics plan for each country, led by the local regulatory contact, and by deciding early whether affected units will be returned, corrected locally or destroyed locally with a certificate. Returning recalled product across a border is a regular export and import, so it needs documents, a classification and an importer of record like any other shipment.
For a cross-border recall, plan for these points:
Local authority requirements. Other regulators have their own rules on field safety corrective actions. In the European Union, the MDCG guidance on vigilance under the Medical Devices Regulation sets out the terms and process. Confirm each country's requirements with the local authorized representative or distributor.
Return or local destruction. Returning units to the United States may be needed for investigation. Otherwise, supervised local destruction with a certificate can be faster and avoids import steps.
Customs documents. A commercial or pro forma invoice that describes the goods as recalled devices returned for investigation or destruction, with quantities, identifiers and value.
US re-entry. Returned devices are imports into the United States and may face FDA review at entry. Agree the entry approach with your customs broker and regulatory team before goods ship.
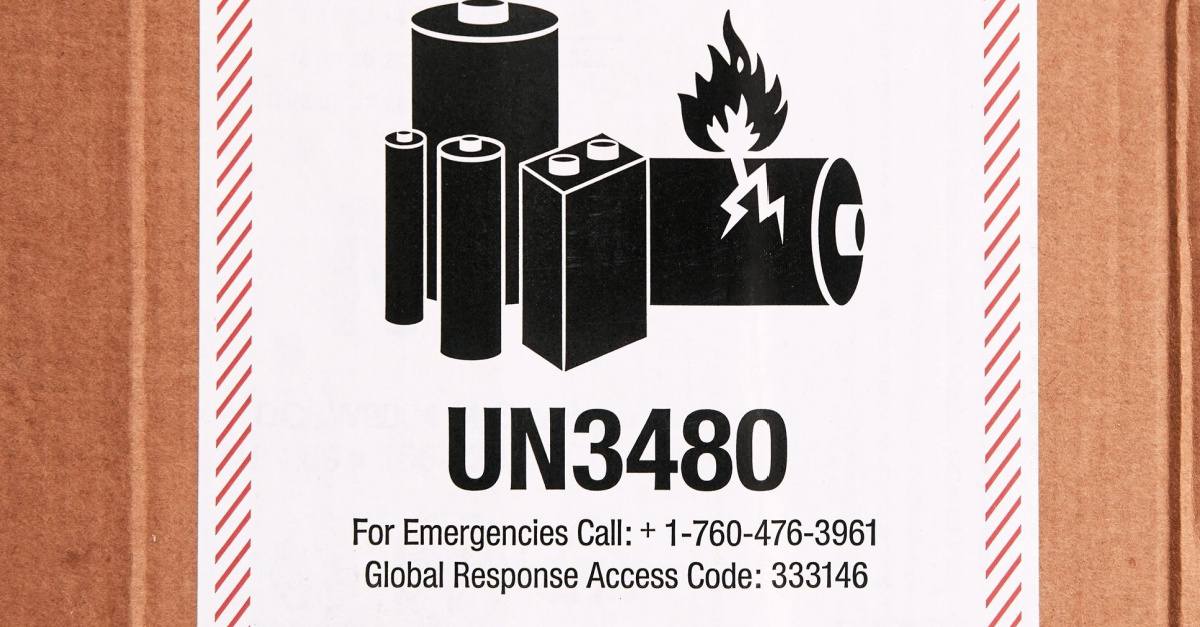
Dangerous goods. Devices with lithium batteries, or returns contaminated with biological material, have their own packing and labeling rules.
Our guide to shipping medical devices internationally and the article on using customs brokers for bio-med shipments explain the customs side.
What records does the FDA expect during a recall?
During a recall, FDA expects records that identify the device and the affected units, list every consignee with quantities and dates, document every recall communication, show the results of effectiveness checks, account for returned or corrected units and describe their disposition. These records feed the Part 806 report, the periodic recall status reports and the request to terminate the recall.
The regulations name these records directly:
Part 806 report. The report must include, among other items, the device identity and UDI or lot and serial numbers, a description of the event and actions, the number of devices subject to the action, the names and addresses of all domestic and foreign consignees with dates and quantities, and copies of all recall communications.
Records of unreported actions. For corrections and removals that are not reportable, 21 CFR 806.20 requires a record with a justification for not reporting, kept for two years beyond the expected life of the device.
Recall status reports. Under 21 CFR 7.53, FDA sets the reporting interval in each case, generally between 2 and 4 weeks. Reports cover consignees notified, responses received, non-responders, quantities returned or corrected, effectiveness check results and the expected completion date.
Termination request. Under 21 CFR 7.55, a firm asks FDA in writing to terminate the recall, with the latest status report and a description of how the recalled product was disposed of.
Logistics generates much of this evidence: pickup confirmations, proof of delivery for recall notices and replacements, receiving logs, temperature records and destruction certificates. Store them against the recall number and the unit identifiers, so the quality team can answer FDA's questions without rebuilding the trail later.
Recall management for regulated health products
Good recall management for regulated health products starts before any recall. Keep distribution records that reach the consignee and lot level, test the process with a mock recall, agree return and replacement procedures with logistics partners in advance and decide who signs off on disposition. The same preparation supports reverse logistics for medical device returns and recalls alike, because the steps for a field return and a recall are close, and only the documentation and urgency differ.
Mercury supports device companies with time-critical courier, air and cold chain services and works alongside their regulatory and quality teams during a recall. For an overview of how Mercury serves this sector, see our medical device logistics page.